
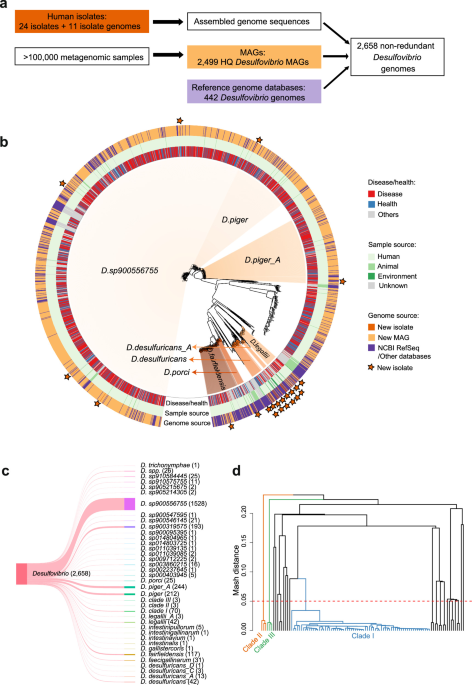
Researchers publishing in Nature have constructed a comprehensive database of 2,658 high-quality Desulfovibrio genomes, revealing extensive genomic diversity and species-specific functional traits associated with diseases such as inflammatory bowel disease and colorectal cancer. The work maps 37 species across 174 metagenomic studies from 32 countries.
Genomic Mapping and Clinical Takeaways
- Database Scale: The curated database incorporates 1,921 new metagenome-assembled genomes recovered from 11,017 human metagenomic samples across 10 cohort studies, alongside 578 genomes from MetaRefSGB and 442 public database entries.
- Species Distribution: Desulfovibrio elongatum sp. nov. (designated D. sp900556755) dominates the dataset, accounting for 57.49% of all analyzed genomes (n = 1,528).
- Functional Divergence: Genetic profiling demonstrates stark species-specific variations in cell motility, urease production, NADH dehydrogenase complexes, antimicrobial resistance genes, and virulence factors.
Genomic Reconstruction Across Global Cohorts
To establish a baseline for characterizing Desulfovibrio spp., the research team applied a metagenome-assembled genome recovery approach to 11,017 samples from 10 diverse human metagenomic cohort studies. These cohorts span varied disease states, ages, and geographies, including Southern China, Japan, Nepal, Tanzania, the Netherlands, the US, Germany, France, and Israel. The majority of the recovered genomes—1,726 out of 1,921—originated from two Dutch cohorts. High prevalence rates appeared in both Dutch studies, while Chinese populations exhibited a wide degree of variation, with maximum prevalence reaching 60% and minimum prevalence dropping to 5%.
After merging sequences from MetaRefSGB and major public repositories like NCBI RefSeq, UHGG, HRGM, and MGnify, the authors de-replicated the entries using a 99.9% average nucleotide identity cutoff. The resulting database spans 37 species from 174 metagenomic studies covering 90 different health conditions. Alongside Desulfovibrio elongatum sp. nov., other major species containing more than 50 genomes include D. piger_A, D. piger, D. sp900319575, and D. fairfieldensis. Culturing efforts yielded 24 newly sequenced human isolates representing eight major species and one uncharacterized lineage, providing crucial material for future functional validation.
Divergent Metabolic Traits and Virulence Profiles
Phylogenetic analysis via pairwise mash distances confirmed that overall genome similarity alone fails to cleanly separate diseased phenotypes from healthy controls, as 686 genomes derived from healthy samples compared to 1,503 from patient samples. Detailed functional profiling, however, illuminated distinct metabolic divisions. Clusters of orthologous groups linked to cell motility showed that a specific flagellin gene (COG1344) is present across 28 species—including D. fairfieldensis, D. desulfuricans, D. legallii, and Desulfovibrio clade III—while remaining entirely absent in D. piger and Desulfovibrio elongatum sp. nov.
| Species / Clade | Dominant Genome Count | Key Genetic Markers | Notable Resistome / Virulence Traits |
|---|---|---|---|
| Desulfovibrio elongatum sp. nov. | 1,528 (57.49%) | Rnf complex (>90%), catB7 gene | High prevalence of chloramphenicol resistance (catB7) |
| D. piger / D. piger_A | Present | RnfCDGEFB gene cluster | Absence of urease operon; catB7 present |
| D. desulfuricans | 42 | ureABCEFH operon, nuo complex | Beta-lactamase resistance (blaDES-1) |
| D. fairfieldensis | 117 | Flagellin gene (COG1344) | Low prevalence of type VI secretion system tssB |
Urease operon genes required for nitrogen metabolism and environmental adaptation appeared in over 95% of genomes for D. desulfuricans, D. legallii, Desulfovibrio sp910584445, and Desulfovibrio sp900547595, yet stayed entirely absent in D. piger. Furthermore, proton-pumping type I NADH dehydrogenase genes (nuoABDHJKLMN) connected to sulfate respiration surfaced in species like D. intestinalis, while an Na+-translocating Rnf complex (rnfCDGEFB) appeared in over 90% of genomes across 18 species.
Antibiotic Resistome Variations in Clinical Isolates
Profiling antimicrobial resistance and virulence factors exposed substantial inter-species divergence. While widespread virulence factors like the elongation factor tufA and catalase katA appeared in over half of all Desulfovibrio species, specialized mechanisms remained restricted.
Antibiotic resistome mapping highlighted targeted resistances across the genus. The chloramphenicol resistance gene catB7 was detected in 211 of 212 D. piger genomes and 1,503 of 1,528 Desulfovibrio elongatum sp. nov. genomes.
Disclaimer: The information provided in this article is for educational and informational purposes only and does not constitute financial advice.
